In 2026, medical consumables manufacturers face a practical challenge: keeping every production step controlled while responding to changing product and market requirements. This guide explores how to ensure compliance in medical consumables manufacturing through clear procedures, documented risk assessments, and consistent quality checks. A batch record, supplier certificate, or training log may seem routine. Each can help explain what happened when a process changes or a concern arises.
The focus is operational, not theoretical. We will look at supplier qualification, material traceability, cleanroom controls, process validation, staff training, and corrective actions. We will also consider how internal audits can reveal gaps before they affect finished products. Quality expert W. Edwards Deming is often associated with this principle: “Quality comes not from inspection, but from improvement of the production process.” It captures why compliance should be built into daily work, not left to final inspection. Still, no checklist guarantees a perfect system. Teams should review evidence, investigate recurring deviations, and adapt controls to the product and applicable requirements. Small details matter. A mislabeled bin or an outdated work instruction can undermine otherwise careful production. The outline ahead offers practical steps, while recognizing that implementation takes ongoing judgment.
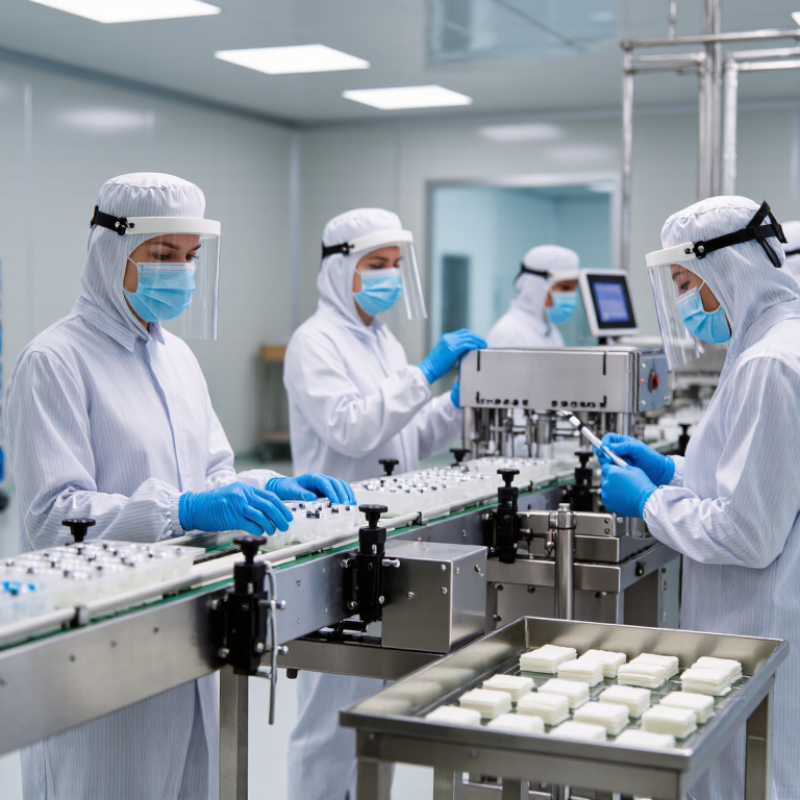
Medical consumables are products routinely used up, replaced, or discarded during care. Examples include gloves, syringes, wound dressings, specimen containers, and infusion tubing. They differ from durable equipment, but “consumable” does not mean low-risk. A sterile dressing touching a wound needs different controls from a non-sterile item used on intact skin.
Compliance begins with a clear intended use and product classification. Requirements vary by market and product, but commonly involve supplier qualification, quality controls, accurate labels, traceable lot and expiry records, and evidence supporting sterility claims. Storage conditions matter too: a crushed sterile pouch or damp carton can undermine confidence in the product.
The WHO’s 2022 Global Report on Infection Prevention and Control estimated that 7 in 100 patients in acute-care hospitals in high-income countries acquire at least one healthcare-associated infection; the estimate was 15 in 100 in low- and middle-income countries. These figures describe infection burden, not cases caused by consumables. Still, they underline why handling and infection-control procedures need attention.
A neat file is not proof of a sound process.
Tips: Check packaging before use, record damaged lots, and review expiry dates during stock rotation. Details matter. When a requirement is unclear, document the question and verify it with a qualified regulatory or quality professional.
Mapping begins with where a product will be sold, what it does, and who uses it. A wound dressing, examination glove, and infusion set may follow different classification routes, even though all are consumables. Record intended use, user population, contact duration, and whether the product is sterile. These details help identify relevant medical-device regulations, registration duties, conformity assessment, and language requirements for labeling. Rules vary by market. Check current guidance from the responsible authorities, not an old distributor checklist.
Next, connect each requirement to evidence. Depending on the product, this may include material specifications, biological safety assessments, sterilization validation, packaging integrity, and shelf-life data. Standards can provide useful test methods, but their status differs across jurisdictions; confirm whether a standard is recognized, adopted, or required for the specific product. Keep a compliance matrix with the requirement, source, version, evidence file, and responsible owner.
Keep evidence close. A spreadsheet can look complete while hiding a weak assumption, so mark uncertain items and revisit them with qualified regulatory or quality staff before making release decisions.
Uncertainty deserves attention.
Qualifying suppliers starts with evidence, not a polished questionnaire. Ask for current quality certificates, audit findings, material specifications, and lot-level traceability. Check whether the supplier can identify the source of a resin or coating and notify you before changing it. Small details matter. A missing revision date on a specification can create confusion months later. During audits, compare written procedures with actual production records, including training logs and inspection results. An audit is only a snapshot, though; follow-up matters.
Materials should be assessed against their intended use and contact conditions. Request composition data, relevant safety information, and test results that support the product’s claims. For items touching skin or tissue, document how material suitability was evaluated. Do not rely on a generic certificate alone. Manufacturing controls need the same scrutiny: review process qualification, equipment maintenance, environmental monitoring where relevant, and acceptance criteria for finished goods. If sterilization or special packaging applies, verify that controls and records cover those steps. Keep changes under review, even when they seem minor. A new supplier sub-tier or adjusted process setting may affect performance. I have seen documentation look complete while leaving one practical question unanswered: who notices when the process drifts? Establish clear owners, escalation steps, and review intervals, then revisit them as product risks change.
| Compliance area | Practical control | Records or evidence to retain | Relevant reference | Review trigger |
|---|---|---|---|---|
| Supplier qualification | Assess suppliers according to the risk and criticality of the supplied material or service. Verify quality-system capability, specifications, change notification, and applicable regulatory responsibilities before approval. | Approved supplier list, qualification questionnaire, risk assessment, quality agreement, evaluation results, and corrective-action records. | ISO 13485:2016, including purchasing controls; applicable national regulatory requirements. | Before initial approval; periodically according to risk; and after significant quality issues, ownership changes, or other material changes. |
| Incoming materials and components | Define acceptance criteria for each item. Use suitable inspection, testing, or supplier documentation to confirm identity and conformity; maintain controls for nonconforming lots. | Current specifications, certificates or test reports where required, receiving inspection results, lot identification, and nonconformance records. | ISO 13485:2016; product-specific requirements and approved purchasing specifications. | For each receipt or lot as specified by the control plan; reassess when specifications, suppliers, or risk change. |
| Material safety and biological evaluation | Evaluate patient-contacting materials in the context of the finished device, intended use, contact type, and duration. Address chemical characterization and biological risks where applicable. | Material composition information, supplier declarations, biological evaluation plan and report, and supporting test or toxicological assessments. | ISO 10993-1:2020 and relevant parts of the ISO 10993 series; applicable market requirements. | When materials, suppliers, manufacturing residues, sterilization, or intended use change, and when new evidence affects the evaluation. |
| Manufacturing process validation | Validate processes when the output cannot be fully verified by subsequent inspection or testing. Establish documented parameters, acceptance criteria, equipment controls, and revalidation rules. | Approved protocols and reports, equipment qualification records, process parameter logs, operator training, and revalidation assessments. | ISO 13485:2016 process-validation requirements; relevant process-specific standards. | Before routine production and after significant equipment, software, facility, material, or process changes. |
| Cleanliness and environmental controls | Set environmental controls based on product and process risk. Define monitoring, cleaning, gowning, contamination-control, and response requirements for relevant production areas. | Environmental monitoring data, cleaning records, maintenance logs, training records, and investigation and corrective-action documentation. | ISO 14644 series for cleanroom classification and monitoring where cleanrooms are used; applicable product and process requirements. | At defined monitoring intervals; after excursions, facility changes, or evidence of contamination risk. |
| Sterilization, when applicable | Select and validate a sterilization method appropriate to the product. Maintain validated cycle or dose controls, routine monitoring, and documented release criteria. | Validation and routine-control records, load or batch records, monitoring results, deviations, and product-release documentation. | ISO 11135 for ethylene oxide; ISO 11137 series for radiation; ISO 17665 for moist heat. Apply the standard relevant to the selected method and current requirements. | Before routine sterilization; at required routine intervals; and after significant changes or process deviations. |
| Packaging and shelf life | Validate packaging that maintains product integrity and, for sterile-barrier systems, sterility until the point of use under specified storage and distribution conditions. | Packaging specifications, validation reports, seal or integrity test results, aging rationale, and distribution simulation records where applicable. | ISO 11607-1 and ISO 11607-2 for packaging of terminally sterilized medical devices. | Before release of a new or changed packaging system; reassess after material, sealing, sterilization, or distribution changes. |
| Traceability and change control | Link relevant production and inspection records to batches or lots. Review proposed changes to materials, suppliers, tooling, software, processes, and specifications before implementation. | Device or batch history records, lot genealogy, change requests and approvals, impact assessments, and required regulatory submission or notification assessments. | ISO 13485:2016 traceability and change-control requirements; applicable UDI and market-specific rules. | For each production batch as defined by the quality system; before implementation of any controlled change. |
| Regulatory quality-system readiness | Maintain a documented quality management system appropriate to the markets served. For products subject to U.S. FDA requirements, account for the Quality Management System Regulation (QMSR), effective February 2, 2026. | Quality manual or equivalent system documentation, internal audit results, management review records, training records, and corrective and preventive action records. | U.S. FDA QMSR (21 CFR Part 820); ISO 13485:2016; EU MDR 2017/745 where applicable. | Through scheduled audits and management review; when entering a new market or when regulatory requirements change. |
Note: Applicability depends on the product, intended use, manufacturing role, and target market. Confirm current regulatory and standards requirements for each product; this table is a planning aid, not a substitute for a product-specific compliance assessment.
Medical consumables compliance depends on controls people can use consistently, not binders that sit untouched. Define acceptance criteria for each product type, including packaging condition, labeling, and any required storage range. At receiving, staff can compare delivery records with purchase details and inspect seals for damage. Small details matter. A clear hold area keeps questionable stock separate while a trained person reviews it.
Documentation should show what happened, when, and who performed the check. Keep current specifications, supplier qualification records, inspection results, training logs, and procedures in a controlled system. For temperature-sensitive items, record readings with dates and device identifiers. If a reading falls outside the approved range, document the assessment and decision before stock is released. Requirements vary by product and market, so quality personnel should confirm which records apply.
Traceability links each item to its source and handling history. Capture lot or batch numbers at receipt, during storage, and at each transfer or issue point. Test the process with a sample: can staff locate affected stock and identify where it went? Keep it practical. Labels may be hard to scan, and records can contain gaps. No checklist catches everything; teams should log near misses, investigate recurring errors, and revise controls when evidence shows a weak point.
Regulatory minimum record-retention periods shown in years
Why it matters: Retaining quality and technical records helps demonstrate that controls were followed and supports product traceability. The U.S. period shown is a floor: records must be retained for the device’s design and expected life when that is longer. EU figures apply to specified technical documentation, declarations of conformity, and certificates.
Sources: U.S. FDA Quality Management System Regulation, 21 CFR 820.35(c), effective February 2, 2026; EU Medical Device Regulation (EU) 2017/745, Article 10(8).
An effective audit follows medical consumables through their actual journey, not just the paperwork. Observe receiving staff checking package integrity, lot numbers, and expiry dates. Compare those records with inventory entries and storage conditions. A thermometer log may look complete while one weekend reading is missing. Small gaps matter.
Use a consistent checklist, but leave room for staff to explain how work happens during busy shifts. Review a sample of records from different days and storage areas. Ask how damaged packaging is identified and separated. A clean storeroom is useful evidence, but it does not prove every item was handled correctly. Audits can miss patterns when reviewers rush.
When a gap appears, record the evidence, risk, owner, and due date. Correct the immediate problem, then look for its cause. If temperature records are repeatedly incomplete, clarify who checks them and what happens when readings are absent. Keep the action practical. Too much paperwork can obscure the fix. Recheck the same process after changes, and document whether the issue returns. If it does, revise the action rather than marking it complete simply because a form was signed.
It is an item routinely used up, replaced, or discarded during care. Examples include gloves, syringes, dressings, and infusion tubing.
Intended use helps determine product classification and suitable controls. A sterile dressing touching a wound needs different safeguards from an item used on intact skin.
Request current quality certificates, audit findings, material specifications, and lot-level traceability. Check revision dates and ask how suppliers report changes.
Review composition data, safety information, and test results against the product’s intended contact conditions. A generic certificate may leave important questions unanswered.
Check process qualification, equipment maintenance, training logs, inspections, and acceptance criteria. An audit is only a snapshot.
Look for crushed pouches, damp cartons, or other packaging damage. Record affected lots and verify expiry dates during stock rotation.
They help teams identify affected products and manage stock rotation. Details matter. Records need to match the items actually received and used.
Assign owners, escalation steps, and review intervals for supplier and process changes. Still, small adjustments can be missed. Revisit controls as risks change.
Ensuring compliance in medical consumables manufacturing starts with clearly defining each product, its intended use, and the quality and safety requirements it must meet. Manufacturers should identify the regulations and product standards that apply to their products and markets, then translate those requirements into practical procedures. A risk-based approach to qualifying suppliers, incoming materials, and manufacturing processes helps prevent quality issues before they affect finished products.
A reliable compliance program also depends on consistent quality controls, accurate documentation, and traceability throughout production and distribution. Clear records make it easier to verify that materials, processes, and products meet established requirements. Regular audits can reveal gaps, while timely corrective actions help address their causes and reduce the chance of recurrence. Together, these practices provide a practical framework for how to ensure compliance in medical consumables manufacturing and support dependable product quality over time.
Ryu Medical